 Materials
science
How to use lammps on ENEA-grid
Application Name:
LAMMPS
Discipline: Molecular modeling
LAMMPS is a classical molecular dynamics code, and an acronym for Large-scale Atomic/Molecular Massively Parallel Simulator. LAMMPS has potentials for solid-state materials (metals, semiconductors) and soft matter (biomolecules, polymers) and coarse-grained or mesoscopic systems. It can be used to model atoms or, more generically, as a parallel particle simulator at the atomic, meso, or continuum scale.
LAMMPS runs on single processors or in parallel using message-passing techniques and a spatial-decomposition of the simulation domain. The code is designed to be easy to modify or extend with new functionality.
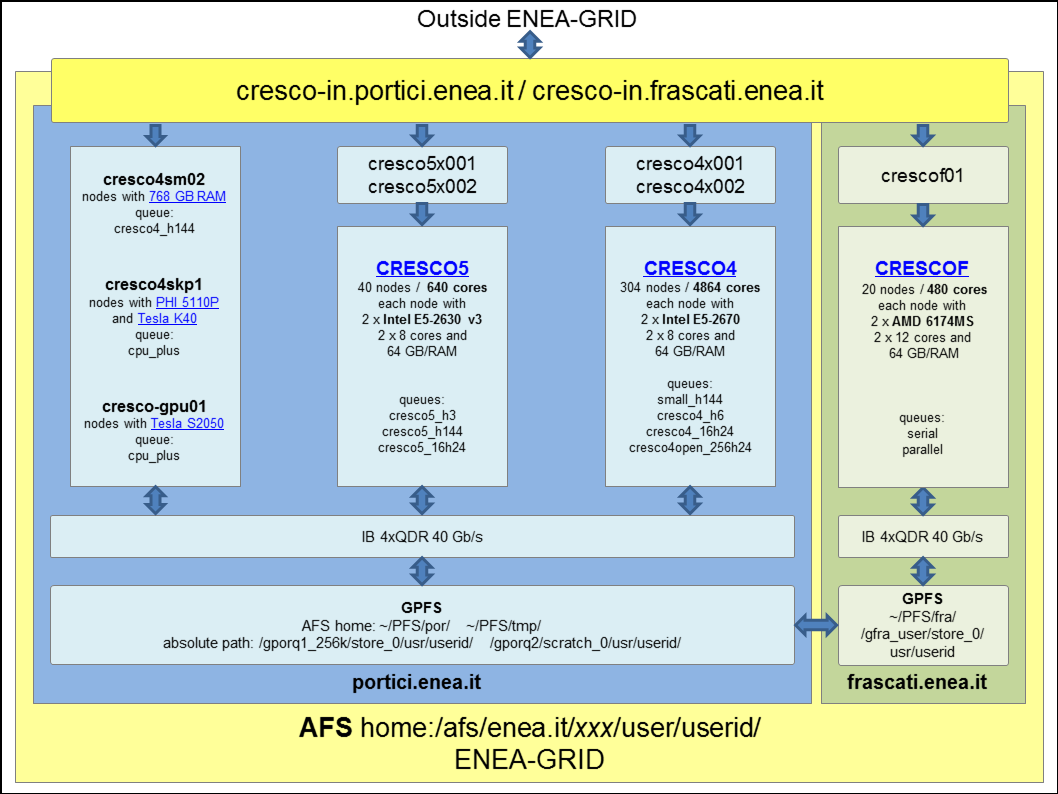
In this page you can find the launching and submission syntax for LAMMPS code on the ENEA-GRID. Please, read the following documentation for running serial/parallel application on CRESCO clusters using LSF framework: Batch Jobs / LSF. This is not a tutorial about using codes like LAMMPS, for that, you may find manual and tutorial on the official web site lammps.sandia.gov.
ENEA-GRID/CRESCO utilization rules (link).
LAMMPS version 31 Mar. 2017 (manual.pdf; manual.html)
bsub -n 400 -q cresco4_256h24 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 64 -q cresco4_16h24 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 64 -q cresco4_h6 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 1 -q small_h144 -i file.in -o file.out -e err.dat lammps17_ser args
example for submission on CRESCO5: cresco5x001, cresco5x002.
bsub -n 128 -q cresco5_16h24 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 1 -q cresco5_h144 -i file.in -o file.out -e err.dat lammps17_ser args
N.B.
Some useful bsub options: -W time_in_minute, -R "span[ptile=16]", -R "span[ptile=24]",
-R "select[hname==crescoxx1 || hname==crescoxx2]", -R "select[hname!=crescoxx3]".
Discipline: Molecular modeling
LAMMPS is a classical molecular dynamics code, and an acronym for Large-scale Atomic/Molecular Massively Parallel Simulator. LAMMPS has potentials for solid-state materials (metals, semiconductors) and soft matter (biomolecules, polymers) and coarse-grained or mesoscopic systems. It can be used to model atoms or, more generically, as a parallel particle simulator at the atomic, meso, or continuum scale.
LAMMPS runs on single processors or in parallel using message-passing techniques and a spatial-decomposition of the simulation domain. The code is designed to be easy to modify or extend with new functionality.
In this page you can find the launching and submission syntax for LAMMPS code on the ENEA-GRID. Please, read the following documentation for running serial/parallel application on CRESCO clusters using LSF framework: Batch Jobs / LSF. This is not a tutorial about using codes like LAMMPS, for that, you may find manual and tutorial on the official web site lammps.sandia.gov.
ENEA-GRID/CRESCO utilization rules (link).
|
CRESCO Cluster |
CRESCO4 front-end cresco4x001 cresco4x002 |
CRESCO5 front-end cresco5x001 cresco5x002 |
||
| LSF submission |
bsub -n N -q queue bsub_options -i file.in -o file.out -e file.err lammps17_par args bsub -n 1 -q queue bsub_options -i file.in -o file.out -e file.err lammps17_ser args |
|||
LAMMPS version 31 Mar. 2017 (manual.pdf; manual.html)
- This version is avaible on CRESCO4 and CRESCO5;
- args are lammps command-line options;
- the following packages are installed.
bsub -n 400 -q cresco4_256h24 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 64 -q cresco4_16h24 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 64 -q cresco4_h6 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 1 -q small_h144 -i file.in -o file.out -e err.dat lammps17_ser args
example for submission on CRESCO5: cresco5x001, cresco5x002.
bsub -n 128 -q cresco5_16h24 -i file.in -o file.out -e err.dat lammps17_par args
bsub -n 1 -q cresco5_h144 -i file.in -o file.out -e err.dat lammps17_ser args
N.B.
- It is not necessary to set the MPI enviroment;
- From each cresco cluster it is possible to use RESTART files generated by the other cresco clusters;
- For the interactive submission on the front-ends cresco4x002 and cresco5x002 use the command:
- lammps17_ser < file.in > file.out
- http://www.ctcms.nist.gov/potentials/
- https://sites.google.com/site/eampotentials/
- https://icme.hpc.msstate.edu/mediawiki/index.php/LAMMPS_tutorials
Some useful bsub options: -W time_in_minute, -R "span[ptile=16]", -R "span[ptile=24]",
-R "select[hname==crescoxx1 || hname==crescoxx2]", -R "select[hname!=crescoxx3]".